
 October 5, 2026
 3 mins to read
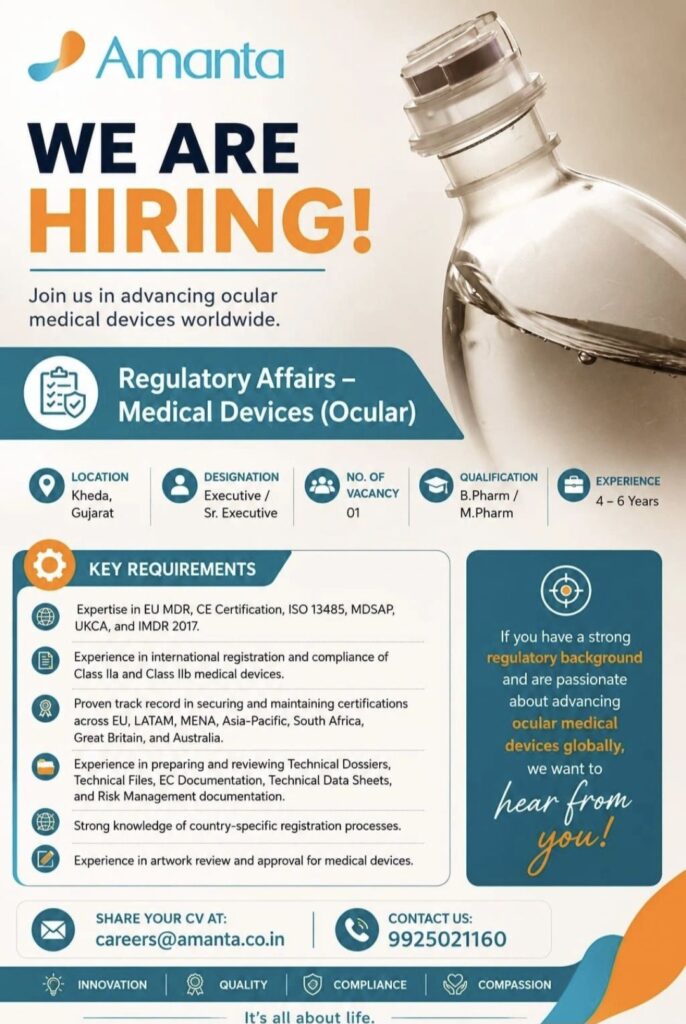
Amanta Healthcare is inviting applications from experienced Regulatory Affairs professionals for its Medical Devices (Ocular) division. This is an excellent opportunity for candidates with expertise in global medical device regulations, product registrations, technical documentation, and compliance management. The position is based in Kheda, Gujarat and offers the chance to work on international regulatory projects involving ocular medical devices.
Job Details
Company Name: Amanta Healthcare
Department: Regulatory Affairs – Medical Devices (Ocular)
Designation: Executive / Sr. Executive
Location: Kheda, Gujarat
Vacancy: 01 Position
Qualification: B.Pharm / M.Pharm
Experience: 4–6 Years
Key Responsibilities
The selected candidate will be responsible for:
- Managing regulatory submissions and registrations for medical devices.
- Preparing and reviewing Technical Files, Technical Dossiers, and Risk Management documentation.
- Ensuring compliance with international regulatory standards and country-specific requirements.
- Coordinating certification activities and maintaining regulatory approvals.
- Reviewing product artwork, labels, and packaging materials for regulatory compliance.
- Supporting global product registrations and post-approval regulatory activities.
Required Skills & Expertise
Candidates should possess strong knowledge and experience in:
- EU MDR (European Medical Device Regulation)
- CE Certification
- ISO 13485 Quality Management System
- MDSAP (Medical Device Single Audit Program)
- UKCA Certification
- IMDR 2017 Regulations
- International registration and compliance for Class IIa and Class IIb Medical Devices
- Regulatory requirements across EU, LATAM, MENA, Asia-Pacific, South Africa, Great Britain, and Australia
- Technical documentation, technical data sheets, and risk management files
- Country-specific registration procedures
- Artwork review and regulatory approval processes
Why Join Amanta?
- Opportunity to work in the rapidly growing ocular medical devices sector.
- Exposure to global regulatory frameworks and international markets.
- Professional growth in regulatory compliance and certification management.
- Collaborative work environment focused on innovation, quality, and compliance.
How to Apply
Interested candidates can share their updated CV to:
Email: careers@amanta.co.in
Technical FAQs
1. What is EU MDR and why is it important for medical devices?
EU MDR (Medical Device Regulation 2017/745) is the regulatory framework governing medical devices in the European Union. It ensures product safety, performance, clinical evaluation, and post-market surveillance before devices can be marketed in Europe.
2. What is the significance of ISO 13485 in medical device manufacturing?
ISO 13485 is an internationally recognized Quality Management System standard specifically designed for medical device manufacturers. It helps organizations maintain consistent product quality and regulatory compliance.
3. What are Class IIa and Class IIb medical devices?
Class IIa and IIb devices are moderate-to-high-risk medical devices. Examples include certain diagnostic, therapeutic, and ophthalmic products. These devices require detailed technical documentation and regulatory approvals before commercialization.
4. What is MDSAP and how does it benefit manufacturers?
The Medical Device Single Audit Program (MDSAP) allows manufacturers to undergo a single regulatory audit that satisfies the requirements of multiple participating countries, reducing audit duplication and improving market access.
5. What documents are included in a Technical File for medical devices?
A Technical File typically includes:
- Device description and specifications
- Design and manufacturing information
- Risk management reports
- Clinical evaluation reports
- Performance testing data
- Labeling and artwork details
- Regulatory declarations and certifications
Leave a Comment